
Research Topics
Dr. Moss’ clinical work focuses on the pathogenesis of cystic lung diseases, such as lymphangioleiomyomatosis (LAM). His research focuses on the post-translational modification of proteins and other molecules by ADP-ribosylation in health and disease.
1. Lymphangioleiomyomatosis (LAM)
The Laboratory of Translational Research has been instrumental in advancing knowledge of LAM, a rare and progressive cystic lung disease that affects women, particularly during their reproductive years. Through a multidisciplinary program that integrates basic science, clinical investigation, and patient-centered research, the laboratory has contributed to defining the diverse clinical manifestations of LAM. The disease is caused by the abnormal proliferation of smooth muscle-like LAM cells, which invade the lung and lymphatic systems, likely through circulating tumor cells. LAM cells are found in nodules lining the lung cysts, which are infiltrated by immune cells and by lymphatic endothelial cells with a mixed blood and lymphatic endothelial cell phenotype. LAM can occur sporadically or as component of tuberous sclerosis complex (TSC), an autosomal-dominant genetic disorder caused by mutations in the TSC1 or TSC2 genes.
LAM is characterized by gradual destruction of lung tissue, with cyst formation. The LAM Protocol helped to define the natural history of LAM, with data on disease progression, clinical variability, long-term outcomes, and patient stratification. Our research has also characterized the molecular and cellular features of LAM cells, which are central to the destruction of lung tissue.
We participated in a multicenter clinical study demonstrating that sirolimus, an mTOR inhibitor, could stabilize lung function in LAM patients (Principal Investigator, Frank X. McCormack). Further research showed that sirolimus therapy could also improve or resolve lymphatic manifestations of LAM, e.g., chylous effusions, lymphangioleiomyomas.
We expanded our research to other destructive lung diseases, such as pulmonary Langerhans cell histiocytosis, Birt-Hogg-Dubé syndrome and idiopathic pulmonary fibrosis (IPF). The success of this research program was made possible by the unique infrastructure and collaborative environment at the NIH Clinical Center.
2. ADP-ribosylation
Dr. Moss and collaborators discovered that the bacterial toxin, cholera toxin, exerted its effects on mammalian cells by ADP-ribosylation, the transfer of an ADP-ribose group from NAD to an acceptor molecule. He later discovered that mammalian cells contain endogenous enzymes with activities that mimic the ADP-ribosyltransferase (ART) activity of cholera toxin. Subsequently, in trying to determine why the ADP-ribosyltransferase activities of mammalian cells are not toxic, Dr. Moss identified ADP-ribosylarginine hydrolases (ARHs), enzymes that release the ADP-ribose moiety from its acceptor. Exquisite control by ARTs and ARHs makes physiological ADP-ribosylation an effective cellular regulatory mechanism with diverse functions.
Dysregulation of ADP-ribosylation is a hallmark of disease. ADP-ribosylation neutralizes the activity of defensin HNP-1, a key effector of the innate immunity. ADP-ribosylated HNP-1 was identified in the airways of patients with pulmonary disease (e.g. asthma, pulmonary fibrosis). HNP-1 is ADP-ribosylated by ART1, one member of the ART family. The ARH family consists of 3 members. ARH1 cleaves ADP-ribose-arginine; mice lacking ARH1 spontaneously develop tumors, indicating a role for ADP-ribosylation in cancer. ARH3 cleaves poly(ADP-ribose) and ADP-ribose-serine. With the Undiagnosed Disease Program (UDP), we identified an ARH3-deficient family, with early death of homozygous individuals. Our transgenic mouse model supported deleterious effects of homozygosity with increased sensitivity to oxidative stress. A potential drug was identified. Thus, transgenic mouse models further elucidate the role of ART and ARH genes in health and disease.
Biography
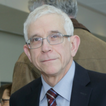
Joel Moss, M.D., Ph.D. is a Senior Investigator in the Critical Care Medicine and Pulmonary Branch, National Heart, Lung, and Blood Institute (NHLBI), National Institutes of Health and Head of the Laboratory of Translational Research and the Lymphangioleiomyomatosis (LAM) program. He graduated Brandeis University, summa cum laude, and received M.D., Ph.D. degrees from New York University School of Medicine. He is a member of Phi Beta Kappa and Alpha Omega Alpha. Following internship and residency (Medicine; Johns Hopkins), he completed post-doctoral and pulmonary fellowships (NHLBI, NIH). He has co-authored over 840 scientific papers, edited/co-authored books, and is co-inventor of patents. Dr. Moss was a member of the NHLBI Institutional Review Board from 1988-2006, its Chair from 1995-2006 and co-authored texts on Ethical Considerations When Preparing a Clinical Protocol (Elsevier, Academic Press, Second Edition). Dr. Moss has received multiple awards, including the Passano Foundation Young Scientist Award, American Foundation for Clinical Research Young Investigator Award, LAM Foundation Awards, the New York University School of Medicine Alumni Achievement Award in Basic Science, the Chiba (Japan) Medical Society Award and the American Thoracic Society Public Advisory Roundtable (PAR) John W. Walsh Award for Excellence. Dr. Moss is an honorary member of the Peruvian Thoracic Society. He is an elected member of the Association of American Physicians, and the American Society for Clinical Investigation (ASCI) and served as ASCI Councilor and Vice-President. Dr. Moss was elected to the Johns Hopkins University Society of Scholars. He is a fellow of the American College of Chest Physicians (FCCP) and the American Thoracic Society (ATSF). Dr. Moss was an Editor of the Journal of Biological Chemistry, Journal of Clinical Investigation, the American Journal of Respiratory Cell and Molecular Biology and the European Respiratory Review. He is currently an Assistant Associate Editor of CHEST and a member of its Commentary Panel. His clinical program focuses on LAM and other cystic lung diseases. LAM results from proliferation of LAM cells and occurs with increased frequency among women with the autosomal-dominant disorder, Tuberous Sclerosis Complex (TSC). Dr. Moss and colleagues initially established the natural history of LAM with detailed clinical phenotyping of LAM and TSC patients and characterized the phenotypic and genotypic features of LAM lung. The team demonstrated that the lymphatic disorders associated LAM were significantly reduced by sirolimus, an inhibitor of the mechanistic target of Rapamycin (mTOR). Dr. Moss has a longstanding interest in ADP-ribosylation pathways. He initially showed that cholera toxin, the pathogenic agent involved in the intestinal disease, Cholera, was an ADP-ribosyltransferase (ART) that uses NAD as an ADP-ribose donor with the amino acid, arginine, and (arginine)proteins as acceptors. Subsequent studies showed that mammalian and avian species have ARTs that catalyze ADP-ribosylation of arginine. Further, enzymes exist that cleave the ADP-ribosyl-arginine(protein) bond (ARHs), regenerating the unmodified protein. The ARH family consist of three members. ARH3 deficiency was found to have early death associated with homozygosity. ADP-ribosylation was shown to modify the activities of regulatory pathways in the heart and lung.
Selected Publications
- Mashimo M, Bu X, Aoyama K, Kato J, Ishiwata-Endo H, Stevens LA, Kasamatsu A, Wolfe LA, Toro C, Adams D, Markello T, Gahl WA, Moss J. PARP1 inhibition alleviates injury in ARH3-deficient mice and human cells. JCI Insight. 2019;4(4).
- Taveira-DaSilva AM, Julien-Williams P, Jones AM, Stylianou M, Moss J. Rates of change in FEV1 and DLCO as potential indicators for mTOR inhibitor therapy in premenopausal lymphangioleiomyomatosis patients. Eur Respir J. 2018;51(4).
- Mashimo M, Kato J, Moss J. ADP-ribosyl-acceptor hydrolase 3 regulates poly (ADP-ribose) degradation and cell death during oxidative stress. Proc Natl Acad Sci U S A. 2013;110(47):18964-9.
- Taveira-DaSilva AM, Hathaway O, Stylianou M, Moss J. Changes in lung function and chylous effusions in patients with lymphangioleiomyomatosis treated with sirolimus. Ann Intern Med. 2011;154(12):797-805, W-292-3.
- Cai X, Pacheco-Rodriguez G, Fan QY, Haughey M, Samsel L, El-Chemaly S, Wu HP, McCoy JP, Steagall WK, Lin JP, Darling TN, Moss J. Phenotypic characterization of disseminated cells with TSC2 loss of heterozygosity in patients with lymphangioleiomyomatosis. Am J Respir Crit Care Med. 2010;182(11):1410-8.
Related Scientific Focus Areas
Molecular Biology and Biochemistry
View additional Principal Investigators in Molecular Biology and Biochemistry
This page was last updated on Thursday, October 8, 2026